



Protein-protein interaction analysis by IP-MS asks which proteins remain with a named bait after antibody or tag capture, negative-control contrast, and LC-MS/MS identification. Selecting a protocol means matching that sentence to a system and a control set, not copying the highest-sensitivity instrument recipe. The analysis fits cells or tissues when an IP-competent antibody exists, or when a tagged handle is treated as AP-MS. The claim is candidate co-recovery, not binary direct binding (Hein et al. 2015). Prefer this selection path when the bait, handle, and negative controls can be named first. Reassess when a Western-only antibody, a missing IgG arm, or a network figure is used to choose the protocol.
Researchers who already have a named bait, handle status, and a written PPI claim can also review the MtoZ Biolabs IP-Mass Spectrometry Service and Protein-Protein Interaction Analysis for project-specific protocol selection.
Select the Experimental System Before the Instrument
IP-MS analysis is a protein-protein interaction method. A protocol is selectable only after the system is named. The system is the bait, the capture handle, and the host background, not the mass spectrometer.
An endogenous bait with an IP-competent antibody is IP-MS. A Flag, HA, or Strep-tagged construct expressed in cells is AP-MS. An immobilized recombinant bait is pull-down MS. Those three handles are compared in IP-MS vs. Co-IP-MS vs. AP-MS: Which Workflow Fits Your Study?. Mixing them in one notebook makes the control set unreadable, so no later instrument setting can repair the file.
The selected system must also match the written PPI sentence. Confirm a named pair, discover unknown partners, or compare the same bait across conditions. A named-pair claim can stay on a tighter wash and a closed blot if a prey antibody works. An open-partner claim needs a digestible eluate and matched IgG through search. A condition comparison must keep antibody, lysis, wash, and elution fixed so the same protein can be compared across arms. Identification depth in a Co-IP-style eluate is typically hundreds to about a thousand proteins, depending on the sample. Depth is not evidence that the selected protocol was better.
The system is a poor first choice when no IP-competent antibody exists and tagging is not allowed, when the sample is a purified domain without a cellular background, or when the claim is reconstituted binding. What a named IP-MS protocol must still answer after this selection is covered in Immunoprecipitation Mass Spectrometry Protocol.
Selecting an IP-MS protocol reports a system that can support candidate co-recovery. It does not name a universally best recipe.
Negative Controls That Travel With the Selected Protocol
Negative controls are part of the protocol, not a later add-on after the bait IP looks clean.
Input is taken before capture. It shows whether the bait existed in that extract and provides a baseline for enrichment. A missing bait protein group is an expression or extraction problem if Input is empty. It is a capture or elution problem if Input is positive. A protocol that changes lysis without a new Input cannot be scored.
IgG or bead controls must use the same lysate batch, lysis, wash, elution, digest, and search. For an antibody IP, IgG is required. Beads without antibody detect resin carryover and do not replace IgG. For a tagged handle, an empty-tag or tag-only arm is required. Empty-tag does not replace IgG on an antibody IP, and IgG does not replace empty-tag on AP-MS. Preclearing is not a substitute for either arm.
If the selected system is a treated or infected lysate, a matched untreated or mock arm must see the same antibody and the same recipe. That arm shows condition-dependent capture. It does not replace IgG.
Selection check |
What a pass looks like |
What the protocol cannot say |
|---|---|---|
Experimental system |
Named bait, stated handle, host background with a protein database |
The mass spectrometer as the protocol |
Negative control |
Input plus IgG or empty-tag on the same recipe |
Every remaining name is bait-specific |
Candidate screening |
Bait recovered, then names ranked versus controls |
An unfiltered search table as the interactome |
Result boundary |
Candidate co-recovery under those arms |
A best protocol or a proven contact |
A practical selection path is:
- Write the PPI sentence the MS table must support.
- Name the bait, handle, and host background.
- Assign Input plus IgG or empty-tag to that same recipe.
- Rank only names that pass that contrast.
- Keep a shortlist for an orthogonal pair assay.
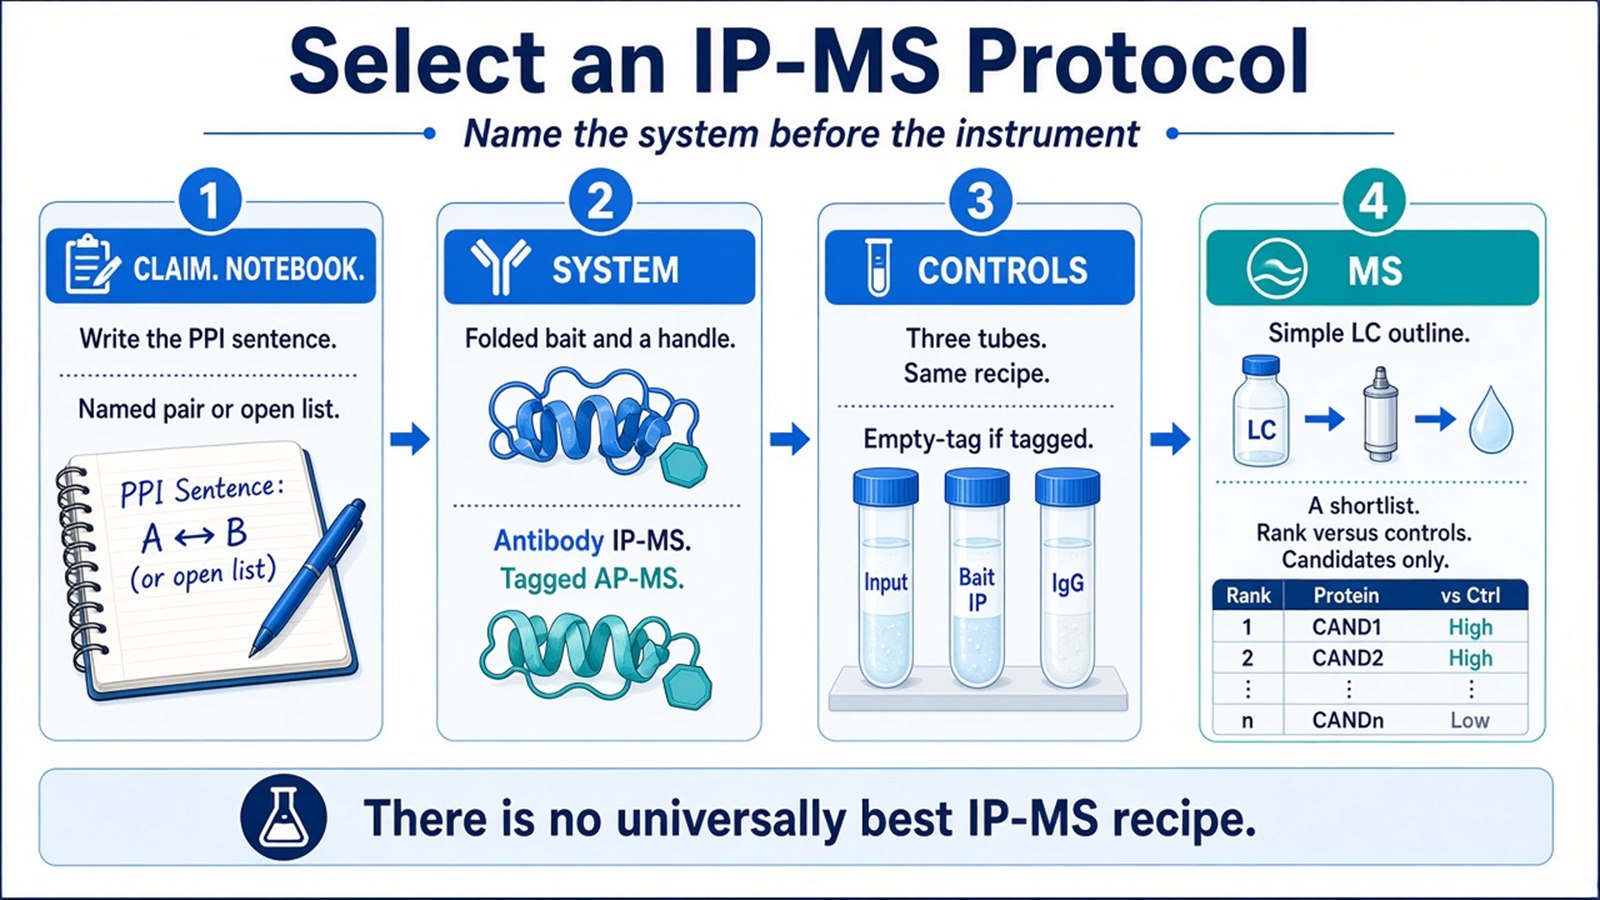
Figure 1. Protocol selection starts after the system is named. A wash cartoon is not a selected IP-MS analysis.
Screening Candidates After the Protocol Is Chosen
Candidate screening starts after bait recovery. If the bait is weak versus Input, stop ranking partners and change handle or extraction before changing the instrument method.
Relative intensity can compare the same protein across Input, IgG or empty-tag, and bait IP only when every arm used one recipe. Intensity does not rank unrelated proteins as tighter binders, and it does not prove that one protocol is more specific than another.
Probabilistic scoring such as SAINT uses bait versus control counts to assign a probability that a prey is specifically co-purified (Choi et al. 2011). A contaminant catalog such as the CRAPome is a prior on control frequency (Mellacheruvu et al. 2013). Pathway software can annotate a shortlist. It cannot repair a missing IgG arm or convert a generic IP cartoon into a validated interactome.
Do not select a protocol because it produced a longer protein list. Do not write a longer list as a more complete PPI network.
Candidate screening ranks names against the controls that were selected with the protocol. It does not certify a binary contact.
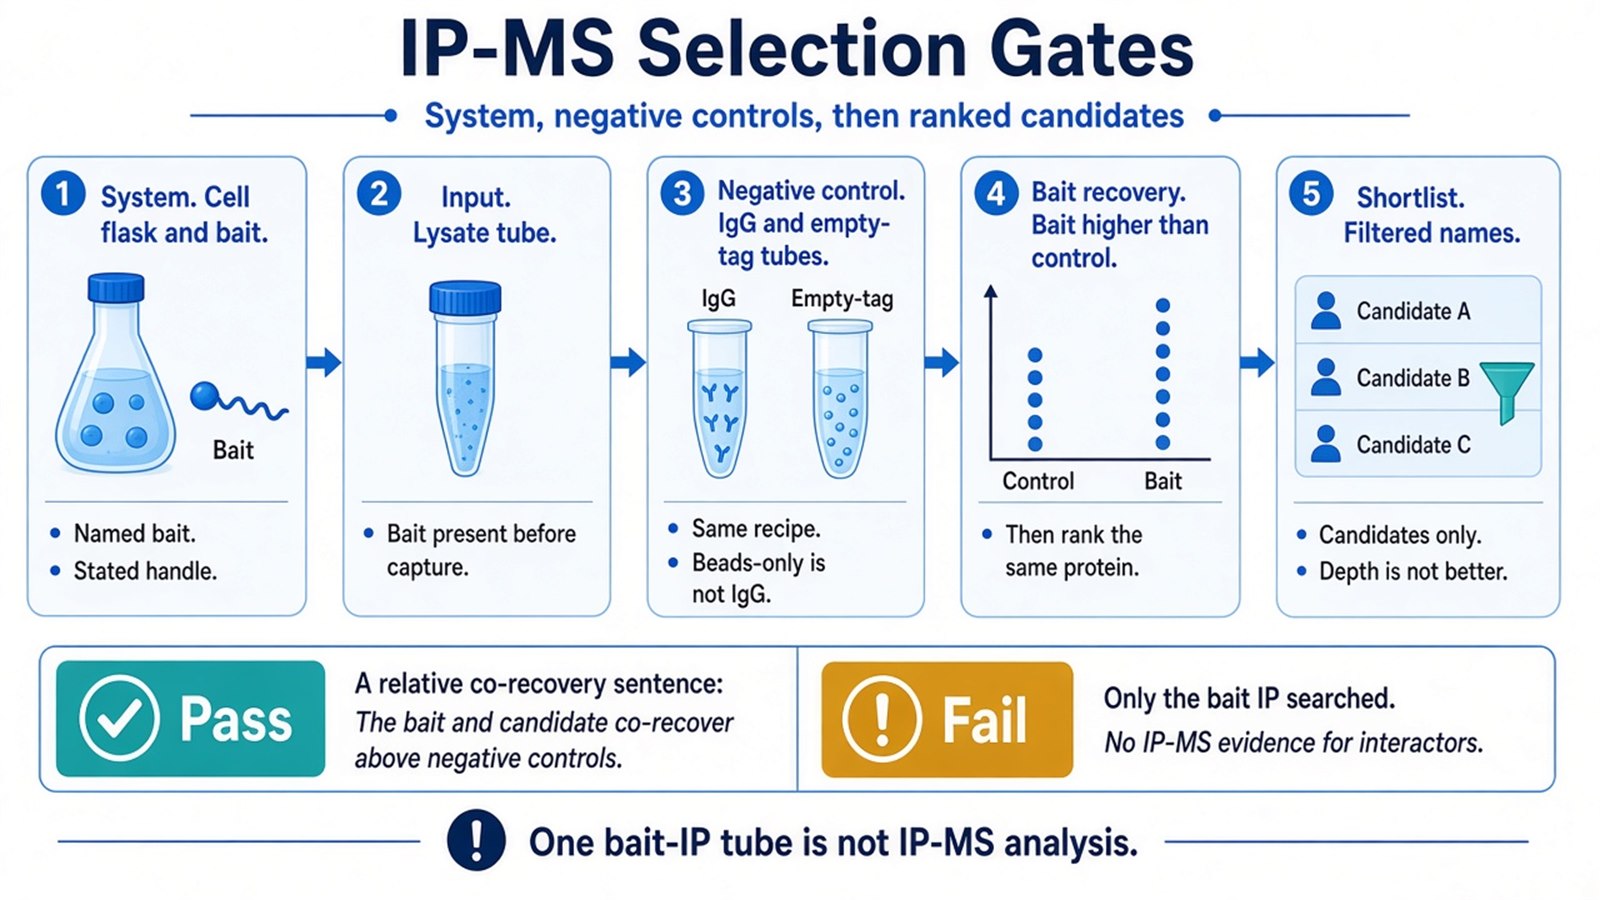
Figure 2. Screening is a control outcome. Extra washing does not replace the selected negative-control set.
Result Boundaries and Common Selection Biases
The common bias is writing IP-MS analysis as highly sensitive, then treating the highest-end instrument or the longest wash as the best protocol. Another bias is sending only the bait IP to MS and reading leftover names as an interaction network. Over-washing can erase the association the claim needed. Under-washing, or a Western elution left in the tube, makes bait IP and controls similar or undigestible.
Do not write a successful bait IP as proof that remaining proteins contact the bait. Do not write a negative table as proof that no interaction exists in the cell. Absence can mean the wrong handle, epitope masking, over-washing, or a bait that is not expressed in the tested state. Do not write antibody IP and tagged AP-MS as interchangeable versions of one best protocol.
Prefer antibody IP-MS when native bait recovery can be shown. Prefer tagged AP-MS when no IP-competent antibody exists. Prefer a defined-pair assay when the next sentence is direct contact.
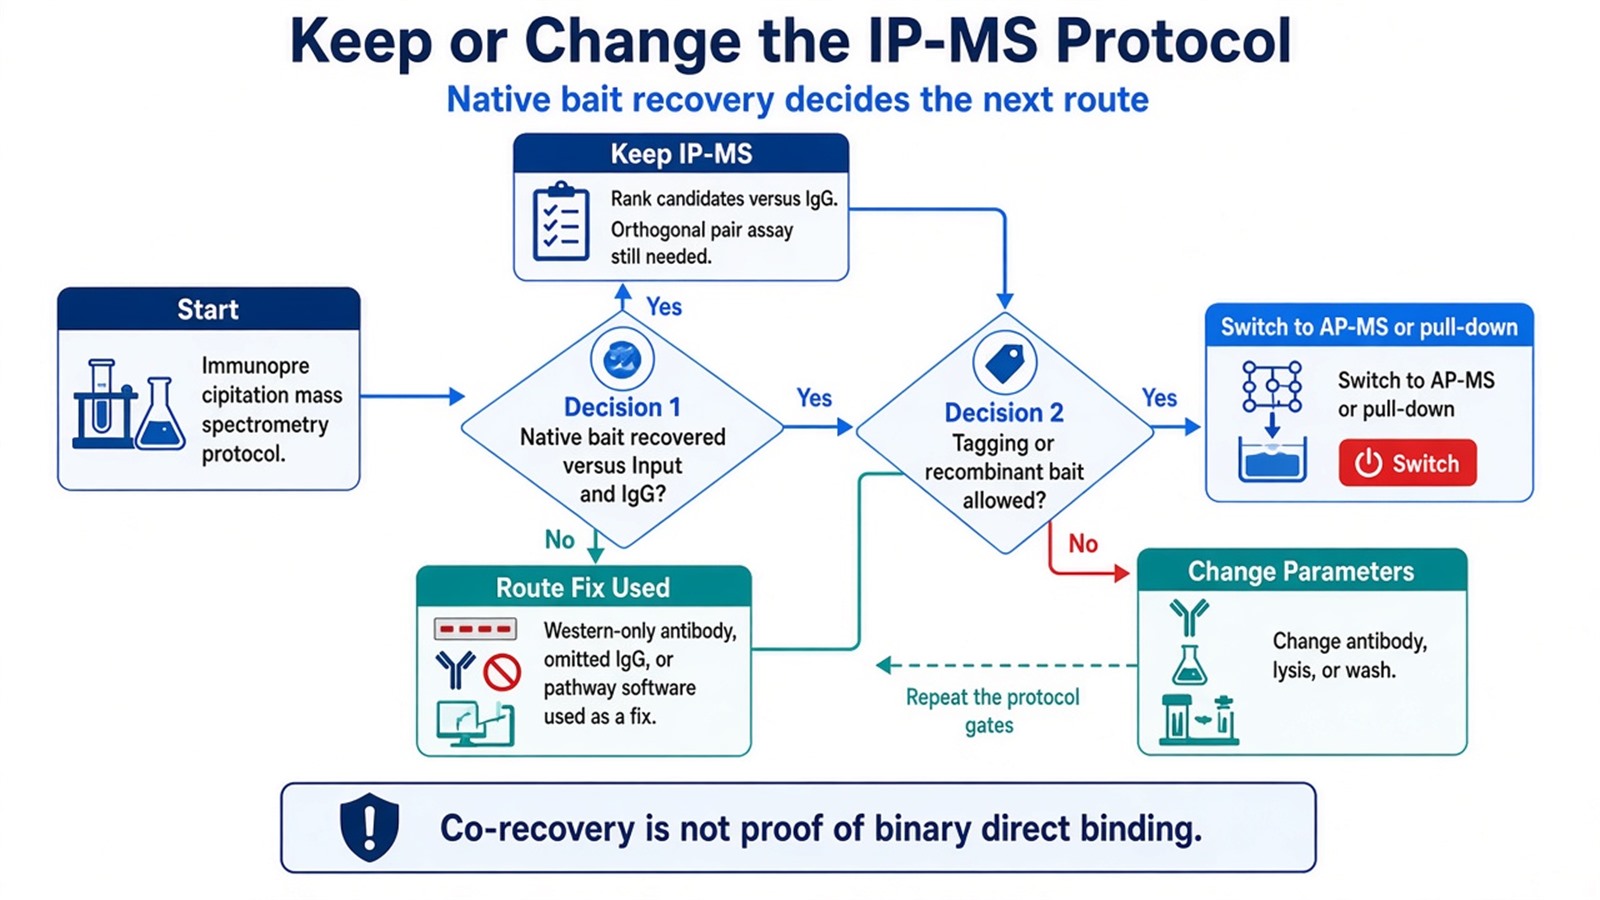
Figure 3. Keep a selected IP-MS protocol only while bait recovery and controls are documented. A pathway map is not a completed selection.
Once the claim, system, handle, negative controls, and the planned digest are defined, the project can be scoped as IP-MS analysis. Researchers can review the MtoZ Biolabs IP-Mass Spectrometry Service for sample evaluation and protocol planning.
Frequently Asked Questions
1. Is there a single best IP-MS protocol?
No. Select the handle and control set that can support the written PPI sentence. A copied wash or instrument method is not a universal answer.
2. Can I select a protocol without IgG?
No. Input and IgG or empty-tag must use the same elution and digest. One bait-IP tube is not an IP-MS result.
3. Is a tagged bait still IP-MS analysis?
No. A tagged construct in cells is AP-MS. Keep empty-tag controls and do not mix that file with an antibody IP.
4. Does a longer protein list mean a better protocol?
No. Depth is typically hundreds to about a thousand proteins, depending on the sample. Depth is not a stronger interaction.
5. When should this selection move to a pair assay?
Move when the next written sentence is direct contact or affinity. Keep the IP-MS shortlist as the input to that assay, not as the proof.
Reference
- M.Y. Hein, N.C. Hubner, I. Poser, J. Cox, N. Nagaraj, Y. Toyoda, I.A. Gak, I. Weisswange, J. Mansfeld, F. Buchholz, et al. (2015). A human interactome in three quantitative dimensions organized by stoichiometries and abundances. Cell, 163, 712-723. https://doi.org/10.1016/j.cell.2015.09.053
- H. Choi, B. Larsen, Z.Y. Lin, A. Breitkreutz, D. Mellacheruvu, D. Fermin, Z.S. Qin, M. Tyers, A.C. Gingras, A.I. Nesvizhskii (2011). SAINT: probabilistic scoring of affinity purification-mass spectrometry data. Nat. Methods, 8, 70-73. https://doi.org/10.1038/nmeth.1541
- D. Mellacheruvu, Z. Wright, A.L. Couzens, J.P. Lambert, N.A. St-Denis, T. Li, Y.V. Miteva, S. Hauri, M.E. Sardiu, T.Y. Low, et al. (2013). The CRAPome: a contaminant repository for affinity purification-mass spectrometry data. Nat. Methods, 10, 730-736. https://doi.org/10.1038/nmeth.2557
