



Relative proteomics answers whether a protein changes between conditions. Absolute proteomics answers how much of a named protein or peptide is present in declared units such as fmol, ng/mL, or ppm. That second question cannot be solved by treating discovery intensities as concentrations. It requires a calibrated strategy: a standard of known amount, a matrix-aware response model, and selective measurement of the surrogate peptide that represents the target.
Many advanced techniques are discussed under the absolute-quantification label. Only some of them truly support calibrated amount reporting. Others remain relative discovery tools, or provide abundance estimates that should not be read as validated concentration. Researchers who need concentration-level reporting for a closed target list can review the MtoZ Biolabs Absolute Quantitative Analysis (AQUA) Service for standard strategy and reporting-unit scoping before labeled peptides are ordered.
What Absolute Quantification Means in Proteomics
An absolute claim reports amount against a known reference. In MS proteomics, that reference is usually a stable isotope-labeled standard matched to a proteotypic peptide, measured together with the endogenous analyte by LC-MS/MS. The light-to-heavy response is converted to concentration through a calibration model established for the reporting range.
This differs from relative methods that compare the same peptide across samples without converting signal into amount units. It also differs from open discovery workflows that survey many proteins without predefining calibrated surrogates. Absolute assays are therefore closed-list methods. They quantify scheduled peptides under declared units; they do not automatically deliver proteome-wide copy numbers for every gene product.
The principle behind peptide-level calibrated reporting is reviewed in Principle of Absolute Quantitative Analysis.
Technique Families Used for Absolute Quantification
Practical absolute work in proteomics usually combines three elements: a labeled-standard strategy, a targeted acquisition platform, and a calibration design that matches the sample matrix. Intensity-based estimators and some labeling chemistries are often mentioned nearby, but they support different claims.
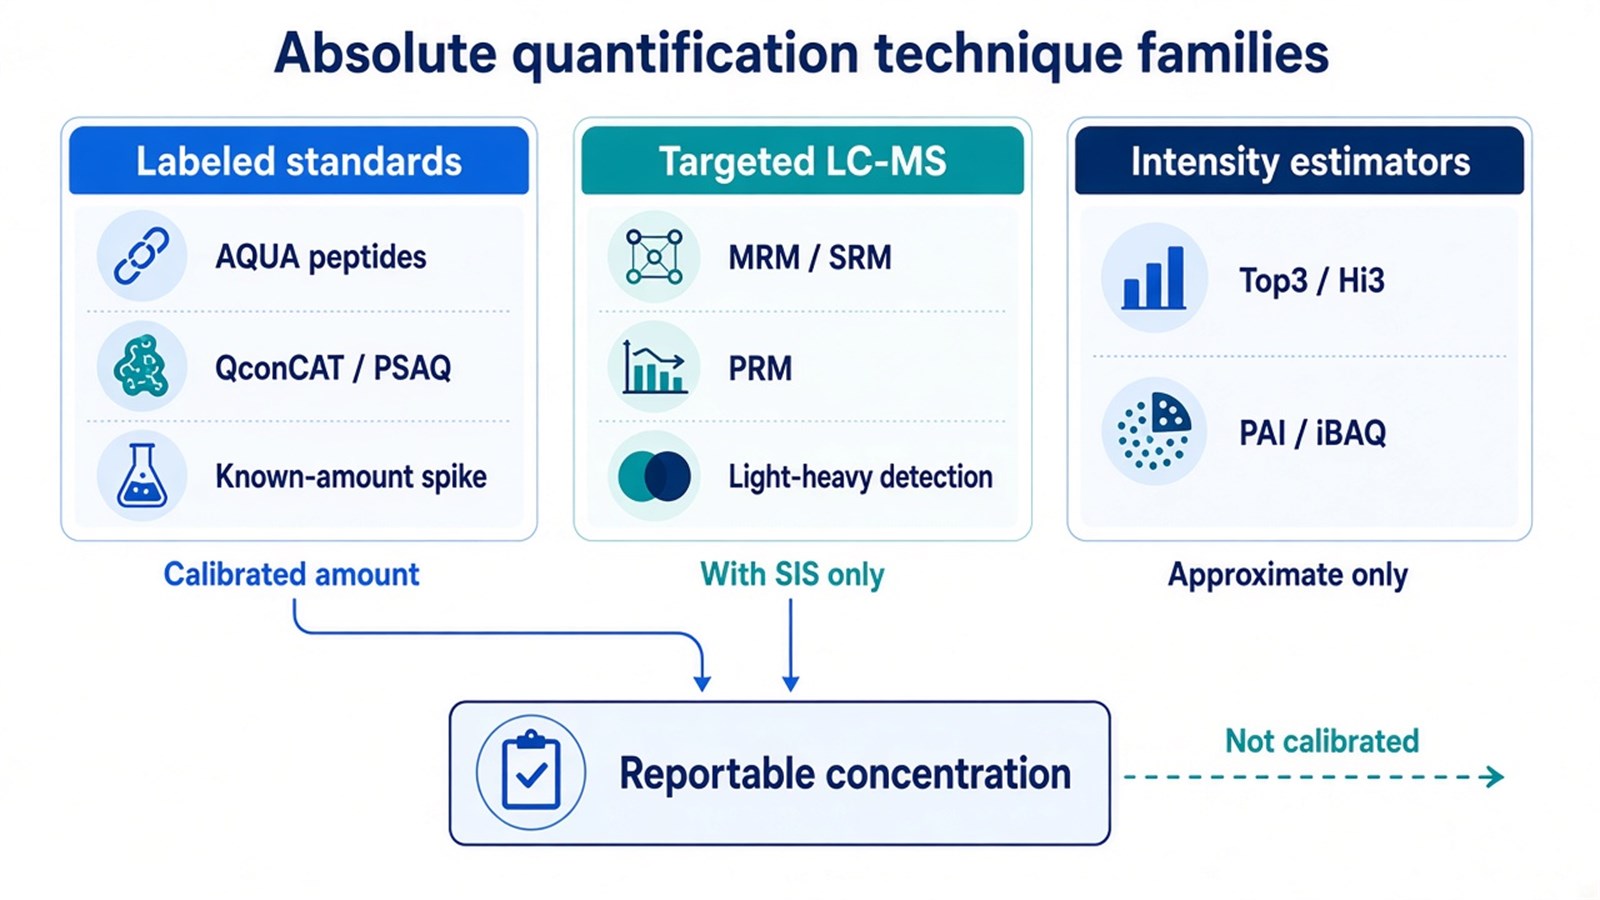
Figure 1. Calibrated absolute reporting depends on labeled standards and targeted detection; intensity estimators remain approximate unless a validated conversion is declared.
1. Stable Isotope-Labeled Peptide Standards (AQUA)
Absolute QUAntification (AQUA) uses synthetic peptides labeled with stable isotopes as internal standards for matching endogenous peptides. The labeled peptide co-elutes closely with the light analyte and is distinguished by mass, so both can be recorded in one LC-MS/MS run. When the standard amount, spike timing, and calibration range are declared, the light-to-heavy ratio supports concentration reporting for that peptide.
AQUA is the most common research route when a short closed panel must be reported in amount units. Peptide-level AQUA does not automatically equal intact-protein copy number if digestion is incomplete, modifications are unaccounted for, or shared peptides confuse protein inference.
2. Protein-Level or Concatenated Standards
Some designs place the labeled reference at the protein or concatenated-peptide level rather than as a single synthetic peptide. Protein Standard Absolute Quantification (PSAQ) uses labeled full-length protein standards. QconCAT constructs concatenate multiple surrogate peptides into one labeled recombinant protein that is digested with the sample. These approaches can better track digestion and preparation losses for selected targets, but they add expression, purification, and characterization burden compared with peptide AQUA.
3. Targeted Platforms: SRM/MRM and PRM
Selected reaction monitoring (SRM), also called multiple reaction monitoring (MRM) on triple-quadrupole platforms, and parallel reaction monitoring (PRM) on high-resolution platforms are the acquisition methods most often paired with absolute standards. They do not create absolute values by themselves. Without a known-amount standard and calibration model, SRM/MRM or PRM remains relative targeted quantitation.
Used with AQUA or related stable isotope standards, these platforms provide the selectivity needed for light-heavy peptide pairs in complex matrices. MRM is often efficient when transitions are stable and interference is manageable. PRM can add fragment-level confirmation when co-eluting background limits MRM performance.
4. Intensity-Based Abundance Estimators
Top3 or Hi3 calculations and Protein Abundance Index (PAI) or related spectral-count indices estimate abundance from discovery MS signals. They can support comparative ranking or approximate stoichiometry when no labeled standard is available. They are not interchangeable with calibrated absolute assays. Without an independent conversion to amount units that has been validated for the project matrix and reporting goal, these estimators should be treated as relative or semi-quantitative readouts.
5. SILAC and iTRAQ in Absolute Context
Stable isotope labeling by amino acids in cell culture (SILAC) and isobaric tags such as iTRAQ are primarily relative quantification chemistries. They compare labeled channels across samples or plexes with high precision for differential analysis. Absolute reporting with these chemistries is possible only when a known-amount reference is introduced and the conversion to concentration is explicitly designed. Labeling alone does not turn a relative experiment into a calibrated absolute assay.
Digital PCR and other DNA absolute methods are outside proteomics absolute quantification and should not be used as substitutes for peptide- or protein-level MS calibration.
How AQUA Becomes a Quantitative Assay
A useful absolute workflow moves from peptide choice to concentration reporting through declared calibration steps.
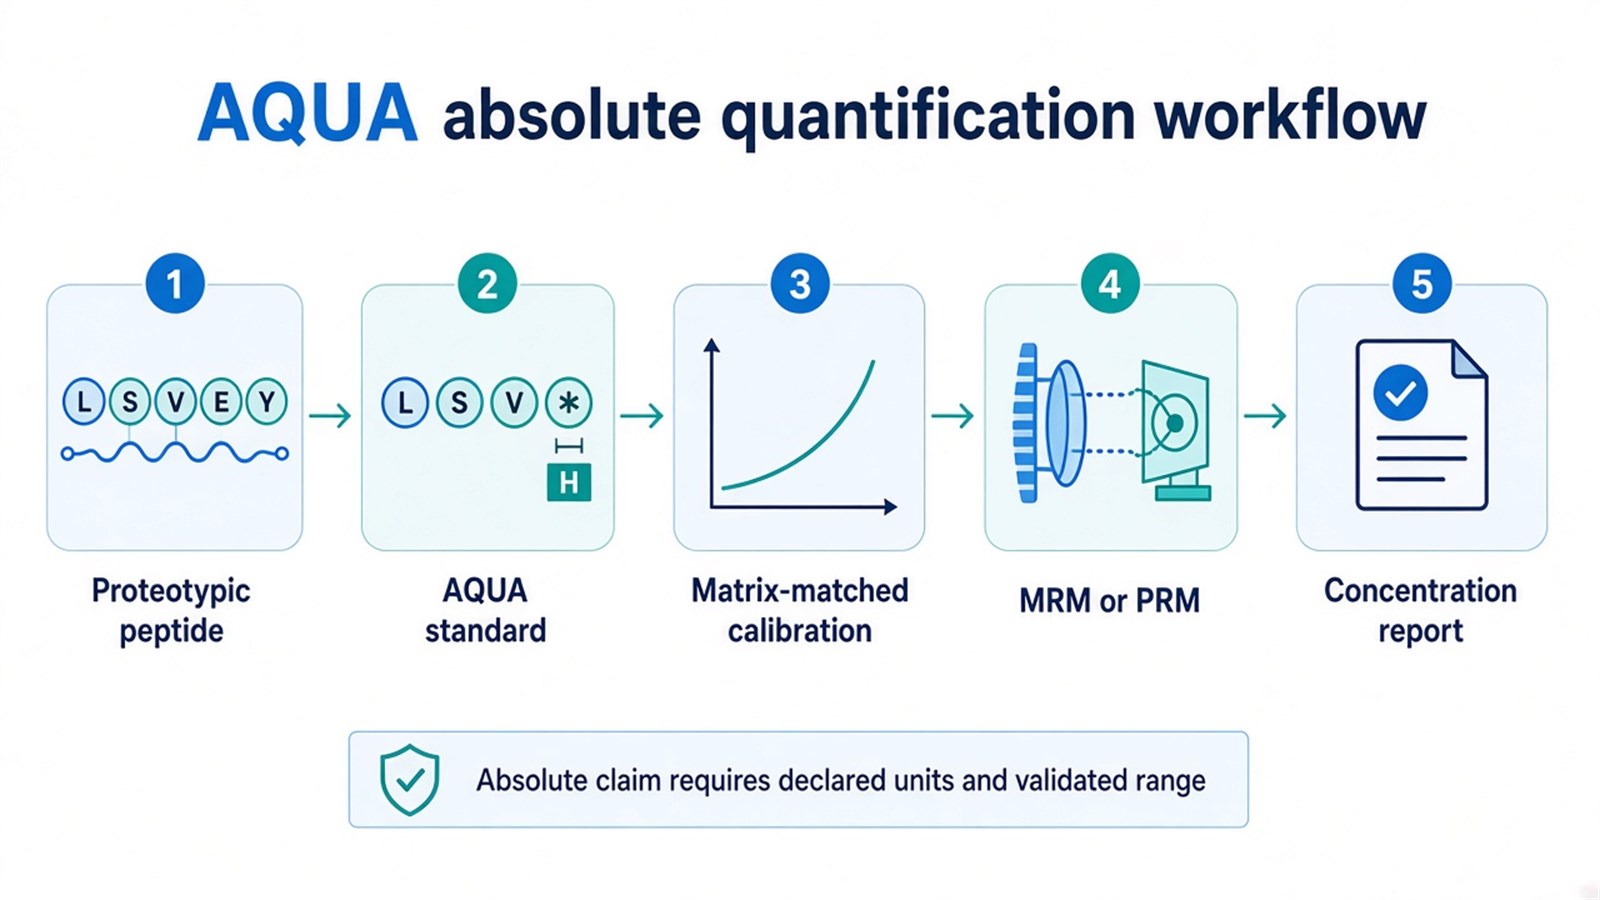
Figure 2. AQUA absolute quantification converts a light-to-heavy LC-MS/MS response into concentration only when calibration and reporting units are declared.
Typical stages include:
- Select proteotypic peptides that represent the intended protein state in the study matrix.
- Design AQUA or related labeled standards and decide spike timing.
- Build matrix-matched calibrators that bracket the expected sample range.
- Acquire light and heavy signals by MRM or PRM.
- Calculate concentration only inside the validated range and report units clearly.
Assay-design choices that often decide whether those stages succeed are covered in Design Absolute Protein Quantification Assay.
How to Choose Among Absolute Techniques
Research need |
Tendency |
Better alternative when |
|---|---|---|
Concentration for a short closed peptide panel |
AQUA with MRM or PRM |
Protein-level PSAQ or QconCAT is required to track digestion recovery more closely |
Multiplex absolute panel with shared digestion control |
QconCAT or multi-peptide AQUA design |
Single-peptide AQUA is enough for one decision target |
Relative discovery or plex comparison first |
SILAC, iTRAQ/TMT, or unlabeled discovery |
Absolute confirmation is already required for named targets |
Rough abundance ranking without standards |
Top3/Hi3 or PAI-style estimators |
Decisions require ng/mL, ppm, or other calibrated units |
Complex matrix with interference risk |
PRM plus labeled standards |
MRM transitions are already clean and validated in matrix |
Sample type, reporting units, and decision use should drive the choice. Plasma biomarker concentration, host cell protein monitoring, and pathway stoichiometry may all use AQUA peptides, yet they differ in matrix handling, calibration depth, and documentation needs.
Practical Limits and Common Misreads
Absolute techniques fail more often at design than at instrument operation. Ordering labeled standards before matrix peptide review, building buffer curves that do not transfer to study samples, or extrapolating beyond calibrators all produce numbers that look absolute but do not support the intended decision. Peptide amount also does not automatically equal protein amount when digestion yield, isoform coverage, or modification state is incomplete.
Diagnostic cutoffs, commercial release decisions, and regulated clinical claims sit outside ordinary research absolute reporting unless a separately validated assay framework is declared.
Conclusion
Advanced absolute quantification in proteomics is not a single technology name. It is a calibrated claim built from labeled standards, selective LC-MS acquisition, and matrix-matched response models. AQUA remains the most accessible peptide-level route for closed panels. PSAQ and QconCAT extend the standard strategy when digestion tracking matters. SRM/MRM and PRM provide the measurement layer only when paired with known-amount standards. Intensity estimators and relative labeling chemistries remain useful nearby tools, but they should not be confused with calibrated concentration reporting.
Researchers planning absolute peptide or protein quantification can contact MtoZ Biolabs to review targets, reporting units, and matrix requirements, and can also consult the MtoZ Biolabs Absolute Quantitative Analysis (AQUA) Service for project-specific feasibility and standard planning.
Reference
- S.A. Gerber, J. Rush, O. Stemman, M.W. Kirschner, S.P. Gygi (2003). Absolute quantification of proteins and phosphoproteins from cell lysates by tandem MS. Proc. Natl. Acad. Sci. U.S.A., 100, 6940-6945. https://doi.org/10.1073/pnas.0832254100
